Formaldehyde: Intersystem Crossing by SOC Surface Hopping
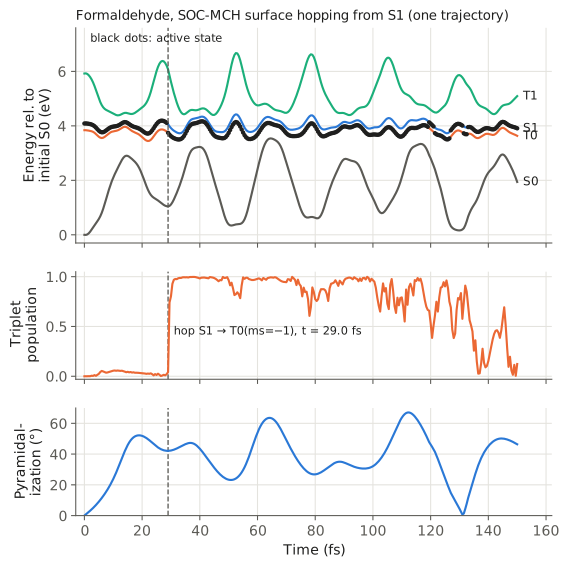
Surface hopping with spin-orbit coupling (SOC-NAMD) lets a trajectory hop between singlet and triplet states. This example starts one trajectory of formaldehyde on S1 (¹nπ*) and follows it for 150 fs. The molecule pyramidalizes, hops to the ³nπ* triplet T0 at 29 fs, and spends most of the run on the triplet before it returns to S1.
Experimental, and needs a build with openqp#535
SOC-NAMD is still under development and must be treated as experimental
(see Ensembles and SOC-NAMD). OpenQP
builds without
openqp#535
stop this run at the first trajectory write after the singlet → triplet
hop, with ValueError: NAMD trajectory schema/model mismatch on append.
The reference trajectory below was run with GitHub main commit 958495f
plus the #535 fix (commit 20272ac5b), not with the build used for the
other pages. Surface hopping is sensitive to rounding, so later hop times
can differ between machines and builds even with the same seed.
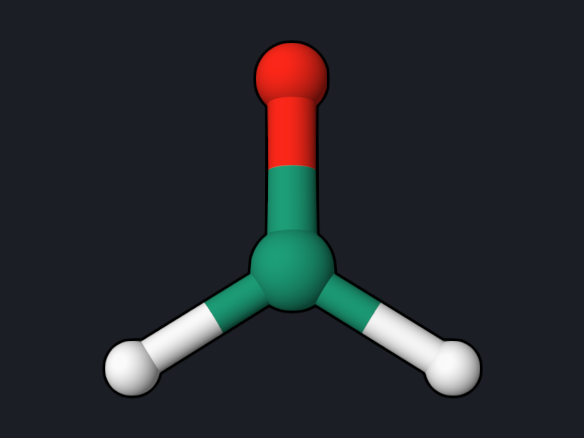
Copy and run
mrsf(nstate=3)/bhhlyp/6-31g* namd(S1,scheme=custom,tdc=npi,rescale=isotropic,thrshe=0.367493,frustrated=reflect,soc=true,soc_basis=mch) md(nstep=300,dt=0.5,temperature=300,seed=3)
geom="""
C      0.000000     0.000000    -0.526342
O      0.000000     0.000000     0.676110
H      0.000000     0.937620    -1.119884
H      0.000000    -0.937620    -1.119884
"""
Download formaldehyde-soc-namd.oqp
openqp formaldehyde-soc-namd.oqp
mrsf(nstate=3)/bhhlyp/6-31g*: three MRSF-TDDFT singlets (S0, S1, S2) and three triplets (T0, T1, T2), the same states as in the SOC example.soc=true,soc_basis=mch: surface hopping among the 12 spin-pure (MCH) states, 3 singlets plus 3 × 3 triplet M_S sublevels, coupled by the SOC matrix elements. The force is the exact gradient of the active MCH state.scheme=custom,tdc=npi,rescale=isotropic,thrshe=0.367493,frustrated=reflect: overlap-based (NPI) time-derivative couplings and isotropic velocity rescaling. The named schemes with analytic couplings support same-spin NAMD only, so SOC-NAMD sets these four choices explicitly (seescheme=custom).md(nstep=300,dt=0.5,temperature=300,seed=3): 300 steps of 0.5 fs, initial velocities drawn from a 300 K Maxwell–Boltzmann distribution with a fixed random seed.
The run takes about 2.5 minutes with 4 threads.
What you get
Each step prints one line. In SOC-MCH runs active gives the MCH basis index
and its state, grad the state whose gradient drives the nuclei, and
pop[S]/pop[T] the total singlet and triplet populations:
   SOC-MCH-NAMD step     57  t=   28.500 fs  active=2:S1  E_tot=-114.26346330  ...  hop=False  ...  grad=S1  pop[S]=0.9792 ...
   SOC-MCH-NAMD step     58  t=   29.000 fs  active=4:T0(ms=-1)  E_tot=-114.26346919  ...  hop=True  ...  grad=T0  pop[S]=0.9580 pop[T]=0.0420
The trajectory file formaldehyde-soc-namd.namd.trj holds all 12 MCH state
energies, complex amplitudes, populations, coordinates, and velocities at every
step. It is read the same way as in the
ethylene NAMD example; here
state_energies has 12 columns in the order S0, S1, S2, T0(−1, 0, +1),
T1(−1, 0, +1), T2(−1, 0, +1).
| Event | Reference trajectory |
|---|---|
| Initial kinetic temperature | 532 K (one random sample of the 300 K distribution) |
| Pyramidalization before the hop | 45° at 25 fs (0° at the start) |
| Hop S1 → T0(M_S = −1) | 29.0 fs, S1–T0 gap 0.31 eV |
| Time on T0 | 97.5 of 150 fs |
| Later hops | T0 → S1 at 119.5, 127.0 and 133.0 fs; S1 → T0 at 121.5 and 131.5 fs |
| Final state | S1 at 150 fs; singlet population 0.88 |
| Total-energy range over 150 fs | 2.1 × 10⁻⁴ hartree (0.006 eV) |
| Energy-conservation check | passed at all 301 steps |
Reading the result
- Pyramidalization comes first. At the planar start the S1–T0 coupling is zero (El-Sayed's rule, see the SOC example), and the triplet population stays below 0.06 for the first 29 fs. S1 formaldehyde is pyramidal, and the out-of-plane angle reaches 45° within 25 fs. The hop to T0 happens at this distorted geometry.
- S1 and T0 move together. Both are nπ* states, so the S1–T0 gap stays between 0.16 and 0.34 eV along the whole trajectory, while S0 oscillates by more than 3 eV with the C–O stretch and the pyramidalization.
- The hop releases the gap as kinetic energy. The isotropic rescaling conserves total energy, and the kinetic energy rises from 0.18 eV to 0.55 eV across the hop step.
- Populations follow the active state after the hop. The triplet
population jumps from 0.04 to above 0.7 within one step. This comes from the
energy-based decoherence correction (
decoherence=edc, the default), which damps the amplitudes of the states that are not active. - One trajectory is not a result. Intersystem-crossing rates and triplet yields need an ensemble of trajectories; see Ensembles and SOC-NAMD.
Try next
- Change
seed=3to other values to run more trajectories; the time of the first hop changes from one trajectory to the next. - Use
soc_basis=adiabaticto propagate spin-adiabatic states. Withseed=3andnstep=80that trajectory hops at 6.5, 23.5 and 38 fs and is on a mostly-triplet state (T0 weight 0.93) after the last hop. Its force uses the weighted-MCH approximation, which is why the md keywords recommendmchfor now.
See also: md keywords, Ensembles and SOC-NAMD, Spin-Orbit Coupling.